Translational Research
Our research program utilizes preclinical models of peripheral T-cell lymphoma to advance our understanding of the mechanisms that drive
lymphomagenesis.
Read Researchlymphomagenesis.
Development of T-cell Lymphoma Mouse Models, Cell Lines and 3D Cellular Systems
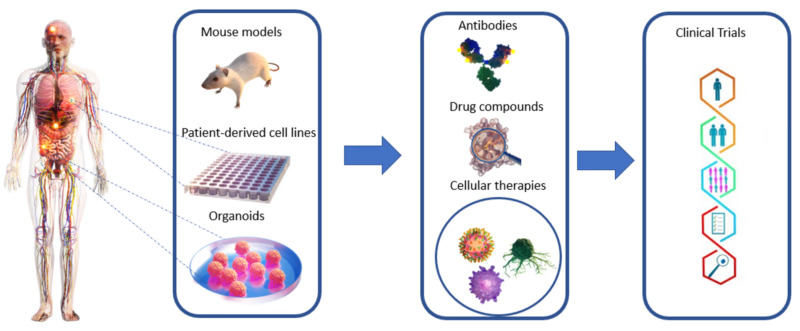
Patient-derived disease models have emerged as critical tools for biological research. Specifically, patient derived xenograft models (PDXs) generally provide higher fidelity representations of the underlying cancer than cell lines. The Jain T-cell lymphoma research team has expertise in generating PDX models and is continually expanding their repository of models representing diverse subtypes of T-cell lymphoma. Cancer cell lines are the most efficient and scalable systems for validating candidates and defining preclinical efficacy. Organoids are the miniature version of in vivo tissues and faithfully recapitulate their architectures. Their co-cultures with immune cells can be leveraged to screen immunotherapeutic compounds for personalized medicine. Through an ongoing collaboration with the Cancer Cell Line Factory at the Broad Institute, we are developing cell lines and organoids for rare types of T-cell lymphoma including hepatosplenic T-cell lymphoma, T-prolymphocytic leukemia, angioimmunoblastic T-cell lymphoma, and enteropathy associated T-cell lymphoma among others.
Detection of Molecular Residual Disease (MRD) in T-cell lymphomas
In aggressive lymphomas, such as diffuse large B-cell lymphoma
(DLBCL), MRD could be prognostic of survival outcomes and
predictive of emerging resistance to therapies. The assessment
of MRD in plasma (where cell-free DNA and exosomes circulate)
is easy to use and could be as informative as cellular DNA. In
addition, NGS technologies are more useful than the classical
"patient allele-specific PCR" because they can identify any
possible clone evolving during the treatment or follow-up, even if
different from that identified at diagnosis, thus forecasting
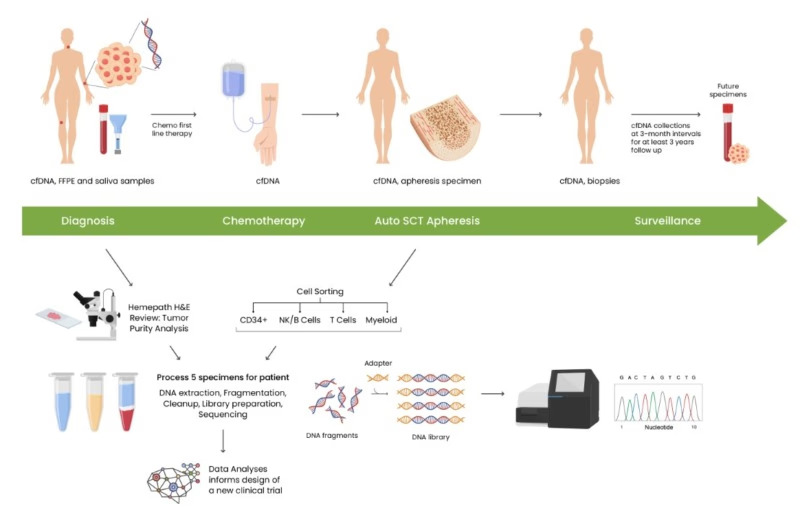
relapse. We aim to use tissue and cfDNA based NGS
approaches to interrogate clonal evolution and relapse in
patients with T-cell lymphomas pre- and post-autologous stem
cell consolidation.
Dissecting the genomic landscape of extranodal natural-killer/T-cell lymphoma (ENKTCL)
The molecular mechanisms underlying resistance to
chemotherapy in extranodal NK/T-cell lymphoma (ENKTCL)
remain poorly understood. To elucidate these mechanisms, we
have assembled a heterogeneous cohort of cases with varying
responses to chemotherapy from multiple centers worldwide,
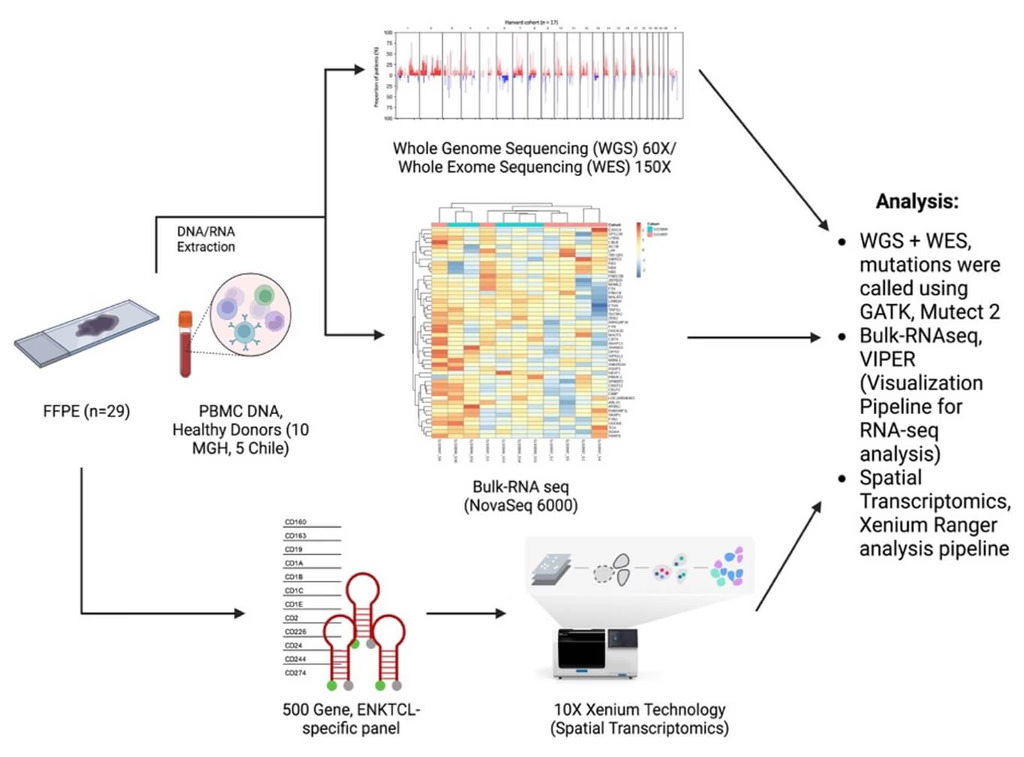
aiming to dissect the genomic landscape of ENKTCL. To
correlate structural variations with somatic mutations, we are
conducting whole-genome, exome, and bulk-RNA sequencing
on diagnostic formalin-fixed paraffin-embedded (FFPE) tumor
tissues, using peripheral blood mononuclear cells (PBMCs) as
healthy controls. In parallel, we have initiated studies to analyze
the spatial transcriptomic and proteomic landscapes of these
cases, employing a custom ENKTCL-specific gene panel with the
Xenium high-throughput in situ analyzer from 10x Genomics. We
aim to utilize this comprehensive multi-omic approach to gain
insights into the unique cell states and pathways contributing to
sensitivity or resistance to chemotherapy, determine spatial
signatures of tumor and tumor-infiltrating immune cells, and
identify biomarkers associated with response and resistance to
therapy in patients with ENKTCL.
The immunological components within tumors, termed the TME,
have long been shown to be strongly related to tumor
development, recurrence, and resistance to therapies. This
remains poorly understood in T-cell lymphomas. We aim to
utilize massively multiplexed single, bulk, and spatial
technologies to untangle the complex relationships between
tumor cells and TME.
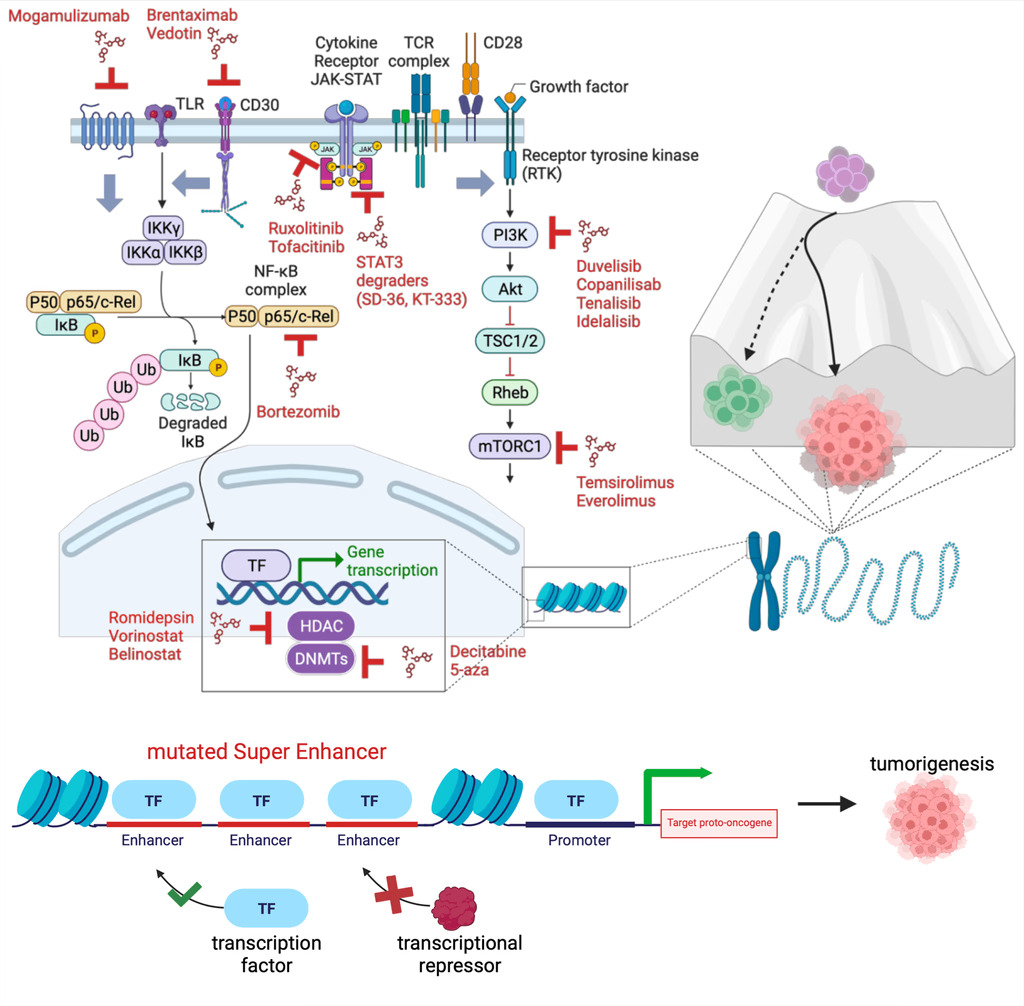
Super-enhancer mutations and functional consequences in T-cell lymphoma
Most NGS-based approaches to advance our knowledge of
PTCL (peripheral T cell lymphoma) biology focus primarily on the
coding regions of the genome, which represent only 2-3% of the
genome. Emerging evidence indicates that non-coding regions
of the genome like enhancers (E) and super-enhancers (SE) can
be recurrently mutated, contributing to tumorigenesis in
lymphomas. The full extent of mutational activity of these non-
coding regions, their specific targets, and their consequent
functional implications remain ill-defined in the most common
PTCL subtypes. By defining these non-coding regions through
multi-omics analysis that centralizes the investigation of 3D
chromatin organization and post-translational histone
modifications in primary tumor samples, cell lines, and patient-
derived xenografts, we can fill the critical gap in our ability to
appreciate the molecular underpinnings of these lymphomas
and, ultimately, precision medicine.